What Is EAEU Pharmaceutical Registration?
The Eurasian Economic Union’s (EAEU) pharmaceutical market represents a significant opportunity for international pharmaceutical manufacturers. The Union brings together Russia, Belarus, Kazakhstan, Armenia, and Kyrgyzstan under a common framework for the circulation of medicines. For manufacturers planning expansion into multiple EAEU markets, the EAEU pharmaceutical registration framework provides a coordinated route for obtaining marketing authorisations across member states.
The reality, however, is more nuanced. The Unified Procedure harmonises important parts of registration, assessment, and pharmaceutical regulation, but it does not eliminate every national requirement. Companies approaching EAEU drug registration therefore need to understand both the common EAEU framework and the requirements that remain relevant at the member-state level.
Planning Pharmaceutical Registration Across the EAEU?
A unified registration framework does not eliminate every national requirement. DDReg Pharma supports pharmaceutical companies with regulatory strategy, dossier planning, and country-specific compliance considerations to help navigate market entry across Eurasian Economic Union member states.
Explore Regulatory Affairs Services →The Architecture of EAEU Pharmaceutical Regulation
The EAEU pharmaceutical regulatory system is based on the Treaty on the Eurasian Economic Union and the Agreement on Common Principles and Rules for the Circulation of Medicinal Products within the EAEU.
The Eurasian Economic Commission (EEC) develops and maintains common regulatory requirements for medicines, including rules for registration and examination, GMP, pharmacovigilance solutions, and other aspects of the medicine lifecycle.
Each member state also maintains its own competent authorities. These authorities participate in the implementation of EAEU requirements and continue to perform national regulatory functions where applicable. The result is a regulatory system that combines Union-level harmonisation with member-state responsibilities.
Each member state retains its own national regulatory authority:
- Russia: Ministry of Health of the Russian Federation, with the Centre for Expertise of Medicinal Products (FEMB/Roszdravnadzor)
- Belarus: Ministry of Health of the Republic of Belarus (Department of Pharmaceutical Inspection)
- Kazakhstan: Ministry of Health of the Republic of Kazakhstan (National Center for Expertise of Medicines)
- Armenia: Scientific Centre of Drug and Medical Technology Expertise (SCDTE)
- Kyrgyzstan: Department of Pharmaceuticals and Medical Devices
These national authorities participate in the Unified Procedure as reference or concerned states, but they retain independent regulatory authority and national registration processes for medicines not submitted through the unified pathway.
EAEU Pharmaceutical Registration Process: Step-by-Step
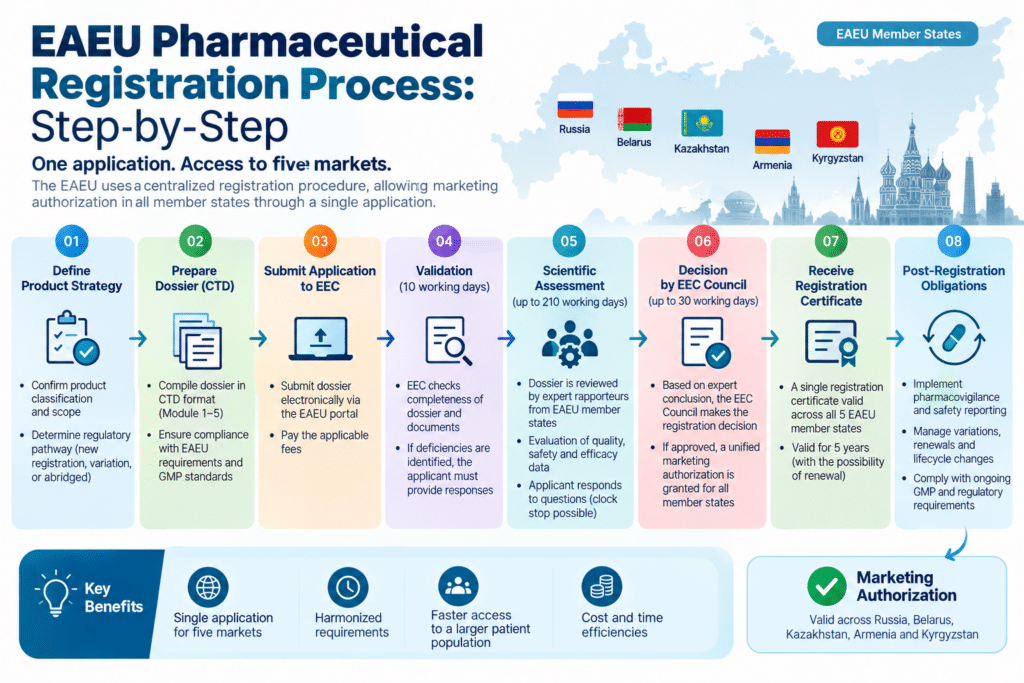
The EAEU registration process generally follows a coordinated sequence:
- Determine the registration pathway: Assess whether the product should be submitted through the EAEU Unified Procedure or a national pathway available under applicable rules.
- Prepare the registration dossier: Compile the product information in the required EAEU CTD format and prepare the applicable national documentation.
- Select the reference state: For the Unified Procedure, the applicant identifies the reference member state responsible for the primary assessment.
- Submit the application: The application and registration dossier are submitted according to the applicable EAEU requirements.
- Scientific assessment: The reference state conducts the main assessment, while other participating states review the assessment and address relevant national aspects.
- Address questions and deficiencies: The applicant responds to regulatory questions and updates the dossier where required.
- Registration decision: Likewise, the title phrase “Unified Procedure” should always be understood as the EAEU procedural framework, not as meaning that all downstream national requirements disappear.
- Post-registration compliance: The marketing authorisation holder must continue to meet applicable requirements for pharmacovigilance, variations, quality, labelling, and other lifecycle obligations.
The process is therefore more than a single submission followed by automatic approval in every market. Regulatory planning needs to account for both the common procedure and member-state requirements.
What Does an EAEU CTD Dossier Include?
The EAEU CTD follows a five-module structure broadly aligned with the international CTD approach:
Module | Main Content |
Module 1 | Administrative and region-specific information |
Module 2 | CTD summaries and scientific overviews |
Module 3 | Quality and CMC information |
Module 4 | Nonclinical study reports |
Module 5 | Clinical study reports |
The registration dossier also needs to include product-specific administrative documents, prescribing information, packaging information, and other supporting documentation required under the applicable EAEU rules.
Language and national documentation requirements should be considered during dossier submission planning rather than treated as a final submission-stage activity. This is particularly important for Module 1 and product information intended for individual member states.
The Unified Registration Procedure: What It Actually Covers
The Unified Procedure allows an applicant to seek marketing authorisation in multiple EAEU member states through a coordinated procedure involving a reference member state and concerned member states.
The reference state conducts the primary scientific assessment and prepares the relevant assessment documentation. The concerned states participate in the procedure and consider the assessment in accordance with the EAEU framework and applicable national requirements.
The procedure therefore creates a common regulatory pathway for multi-state registration, but it does not mean that every aspect of market entry is identical across the five countries.
One EAEU Procedure. Different National Considerations.
Pharmaceutical companies must assess how the unified EAEU framework applies to their target markets, including any relevant national procedures, implementation timelines, and local requirements. Build a registration strategy that accounts for these differences before planning your launch.
Discuss Your Registration Strategy →EAEU Eligibility for the Unified Procedure
The Unified Procedure is available for new medicines seeking registration in two or more member states simultaneously. Certain categories of products, including innovative medicines, orphan medicines, and medicines for serious or life-threatening conditions, are designated for the unified pathway. Generic medicines can also use the Unified Procedure.
However, the Unified Procedure is not mandatory for all products. Applicants may choose to pursue national registrations in individual member states where the commercial priority does not justify a multi-state filing. For Russia, which represents the dominant share of the EAEU pharmaceutical market value, many applicants file nationally rather than through the unified pathway, particularly for time-sensitive launches.
EAEU Unified Registration vs National Registration: What's the Difference?
The choice between unified and national registration depends on the product, target markets, commercial strategy, and applicable regulatory pathway.
EAEU Unified Registration | National Registration |
Designed for registration in multiple EAEU member states | Focuses on an individual member state |
Uses the EAEU common regulatory framework | Follows the applicable national framework |
Involves a reference state and participating concerned states | Assessment is conducted through the national authority |
Useful when multiple EAEU markets are part of the launch strategy | May be relevant when only one market is being targeted or a national pathway applies |
A unified procedure can provide regulatory coordination across markets, while a national pathway may be appropriate when the commercial objective is limited to a specific member state.
EAEU MRP vs DCP: What's the Difference?
The EAEU Unified Procedure is sometimes compared with the EU Mutual Recognition Procedure (MRP) or Decentralised Procedure (DCP), but the systems should not be treated as equivalent.
In the EU:
- MRP is generally used when a medicinal product already has a marketing authorisation in one member state and that authorisation is subsequently recognised in other member states.
- DCP allows an applicant to seek authorisation simultaneously in several EU member states when the product has not yet been authorised in an EU member state.
The EAEU Unified Procedure has its own legal and institutional framework. It involves a reference member state and other participating states, making its structure closer in concept to a coordinated multi-state procedure than to EU MRP. However, it should not be described as an EAEU version of either EU MRP or DCP. Understanding this distinction is important when planning a regulatory affairs strategy for both EAEU and EU markets.
What the Unified Procedure Does Not Unify
The term “unified” is accurate in a procedural sense, but it does not mean that every downstream regulatory requirement is identical across the Union.
- National Pricing and Reimbursement – Marketing authorisation does not create a single EAEU pricing or reimbursement system. Pricing, reimbursement, formulary access, and related health-system decisions remain subject to the relevant national frameworks. Companies therefore need to plan market access activities separately from the regulatory registration itself.
- Labelling and Language Requirements – EAEU requirements provide a harmonised regulatory framework for product information, but language and certain national documentation requirements still need to be addressed for individual markets. Applicants should plan translations, product information, packaging materials, and applicable national requirements early in the registration process.
- EAEU Pharmacovigilance Requirements – The EAEU has established common Good Pharmacovigilance Practice requirements, while member states maintain national pharmacovigilance systems under the Union framework and their applicable legislation.
For marketing authorisation holders, this means that an EAEU registration does not remove the need to understand national reporting channels, competent authorities, contacts, and implementation requirements.
The EAEU Agreement specifically requires member states to maintain effective national pharmacovigilance systems in accordance with EAEU GVP requirements and national legislation. A unified marketing authorisation therefore does not mean that post-authorisation safety obligations can be managed without considering the individual member states.
EAEU vs EU Pharmaceutical Registration: What Are the Key Differences?
The EAEU and EU both use regional regulatory frameworks to facilitate access to multiple markets, but their legal structures and procedures are different.
EAEU | EU |
Five member states: Russia, Belarus, Kazakhstan, Armenia, and Kyrgyzstan | 27 EU member states |
Common EAEU pharmaceutical rules developed through the EEC | EU pharmaceutical legislation and EMA/member-state systems |
Unified Procedure uses a reference state and participating states | Centralised, MRP, DCP, and national procedures are available depending on the product |
EAEU CTD requirements apply within the Union framework | EU submissions use EU requirements and applicable eCTD standards |
National requirements continue to matter for certain post-authorisation and market-access activities | National requirements also remain relevant, particularly for non-centralised procedures and market access |
For companies expanding across both regions, an existing EU dossier can provide a useful starting point, but it should not be assumed to be directly transferable without EAEU-specific assessment and adaptation.
Russia's National Registration Pathway in EAEU
Russia remains an important pharmaceutical market within the EAEU. Products submitted through a national Russian pathway are assessed under Russian legislation and procedures rather than the EAEU Unified Procedure.
Companies considering Russia separately should therefore assess the applicable national registration requirements, dossier expectations, language requirements, clinical evidence, manufacturing compliance, and post-registration obligations before selecting the pathway.
Russian Dossier Requirements in EAEU
Russia accepts CTD-based dossier structures, but EAEU and Russian requirements may require specific adaptations to administrative documentation, product information, language, and other submission elements.
The extent of Russian-specific requirements depends on the product, application type, and applicable regulatory pathway. These requirements should be assessed during regulatory strategy and dossier planning rather than assumed from an existing EU or US submission.
EAEU Pharmacovigilance: What Marketing Authorisation Holders Need to Plan
Pharmacovigilance remains an important part of the EAEU regulatory lifecycle. The EAEU has established common GVP requirements covering areas such as safety information, individual case safety reports, signal management, risk management, and pharmacovigilance system responsibilities.
At the same time, marketing authorisation holders need to consider how these requirements are implemented through the national pharmacovigilance systems of the member states.
A practical EAEU pharmacovigilance strategy should therefore address:
- Applicable EAEU GVP requirements.
- National competent authorities and reporting mechanisms.
- Individual case safety reporting requirements.
- Signal and risk management activities.
- Pharmacovigilance system documentation and oversight.
- Ongoing post-authorisation safety obligations.
Strategic Considerations for EAEU Market Entry
- Choose the registration pathway based on the target markets.
If several EAEU markets are part of the launch strategy, the Unified Procedure can provide coordinated regulatory assessment. If the commercial objective is limited to one country, a national pathway may be relevant where permitted. - Build the EAEU CTD around the applicable requirements from the start.
Adapting an existing global dossier without assessing EAEU-specific requirements can create avoidable deficiencies. - Plan language and product information early.
Translation and national product-information requirements should be incorporated into the submission plan rather than addressed immediately before filing. - Prepare pharmacovigilance arrangements before approval.
Understanding EAEU GVP requirements and their national implementation helps ensure that the safety system is ready when the product enters the market. - Separate regulatory approval from market access planning.
Marketing authorisation is an important step, but pricing, reimbursement, supply, and other commercial requirements may follow separate national processes.
Are Your CMC Documents Aligned with EAEU Requirements?
Quality documentation must be consistent, complete, and aligned with applicable registration requirements. DDReg Pharma provides CMC regulatory support to help assess technical documentation, identify potential gaps, and prepare pharmaceutical dossiers for regulatory submission.
Get CMC Regulatory Support →What Changed in EAEU Pharmaceutical Registration After December 2025?
The transition period for aligning nationally registered medicines with Eurasian Economic Union (EAEU) requirements reached its general deadline on 31 December 2025. Key implications include:
Dossier Alignment: Companies must verify whether required alignment applications were submitted by the deadline.
Certificate Validity: Certain national certificates may remain valid under specific transitional provisions.
Country-Specific Rules: Russia may apply different transitional measures from other EAEU member states.
Regulatory Compliance: Companies should review registration status, outstanding submissions and applicable deadlines.
Market Continuity: Product-by-product assessment is essential to avoid potential regulatory and supply disruptions.
The deadline did not automatically invalidate every national registration certificate; applicable conditions must be checked individually.
Labelling and Language Requirements
Each member state requires product labelling in its official language(s). In the Unified Procedure context, this means the applicant must prepare and validate labelling in Russian (required in Russia, Belarus, and Kazakhstan), Armenian (required in Armenia), and Kyrgyz and/or Russian (required in Kyrgyzstan).
The label content requirements beyond language also contain member-state-specific variations in required statement s, storage condition formatting, and the inclusion of local regulatory contact information. EEC has produced harmonised label guidelines, but national implementing rules add locally specific requirements that the harmonised guidelines do not fully resolve.
Pharmacovigilance Obligations Remain Nationally Fragmented
The EAEU has been working toward harmonised pharmacovigilance requirements, and the EEC has issued EAEU GVP standards. However, the practical pharmacovigilance obligations that MAHs face across the Union remain substantially nationally differentiated.
Russia requires adverse event reports to be submitted to Roszdravnadzor’s national pharmacovigilance database. The reporting timelines, case format requirements, and periodic safety update submission schedules differ between Russia and other member states. Holders of EAEU-unified marketing authorisations are responsible for maintaining pharmacovigilance compliance in each member state independently, with no single reporting pathway that satisfies all national obligations simultaneously.
A unified marketing authorisation does not mean a unified pharmacovigilance system. Companies that assume EAEU registration automatically handles their PV obligations across member states will find themselves non-compliant in one or more jurisdictions within the first post-registration reporting cycle.
Russia's National Registration Pathway
For most pharmaceutical companies, the Russian national market is the commercial objective that justifies EAEU registration investment. Russia’s national registration pathway — for products not submitted through the Unified Procedure — operates under the Federal Law on the Circulation of Medicines (No. 61-FZ) and involves submission to the Ministry of Health through the Gosudarstvennyi Reestr Lekarstvennykh Sredstv (GRLS) system.
Russian Dossier Requirements (CTD with Local Adaptations)
Russia accepts CTD-format dossiers but requires significant Russian-language documentation. The clinical overview, summary of clinical efficacy, summary of clinical safety, and other Module 2 documents must be submitted in Russian. Full translation of the clinical study reports (Module 5) is generally not required for innovative products relying on foreign clinical data, but Russian-language summaries must meet specific content standards.
The Russian Ministry of Health requires that clinical efficacy data include representation of studies in Russian patients or, where this is absent, scientific justification for extrapolation of foreign trial data to the Russian population. For products studied predominantly in Western European or North American populations, this can generate significant procedural questions.
Historical Requirement and Its Current Status of Local Clinical Trials
Russia historically required local clinical trials as a condition of registration for innovative medicines, a requirement that was a major barrier for international manufacturers. The requirement for mandatory Russian Phase III clinical trials has been substantially relaxed in recent years, with the Ministry of Health implementing risk-based approaches to local study requirements. For medicines approved by ICH-member agencies (US – FDA, EMA, PMDA) and supported by robust multinational trial data, full-scale local trials are now rarely required. Post-registration safety and effectiveness studies in Russian patients, however, remain a common condition attached to initial marketing authorisation.
Registration Timelines in Russia
Russia’s target registration timeline for innovative medicines is 160 working days from acceptance to decision. In practice, the timeline is often longer, particularly for products where clinical or CMC questions are raised. Expert review by the FEMB is substantive, and questions can generate multiple review cycles. A total elapsed time from submission to registration decision of 12 to 24 months is common for new products.
Strategic Recommendations for EAEU Market Entry
- Assess commercial priority before choosing between Unified Procedure and national registration.
If Russia is the primary target, a national Russian registration may be faster and more controllable than waiting for the EAEU unified procedure timelines. If simultaneous launch across multiple member states is a strategic priority, the Unified Procedure adds regulatory coordination value despite its extended timelines. - Invest in Russian-language regulatory capability.
All formal submissions to Russian, Belarusian, and Kazakhstani authorities require Russian-language documentation. Applicants who rely on machine translation of clinical overviews and safety summaries will generate avoidable deficiencies. Professional regulatory-grade translation by subject matter experts is not optional. - Plan pharmacovigilance infrastructure before registration, not after.
Each member state requires separate adverse event reporting mechanisms, separate contacts, and (in Russia) registration in the national pharmacovigilance database. Companies that assume post-registration setup is feasible will be non-compliant from day one of market authorisation. - Engage with pricing registration in parallel with marketing authorisation proceedings.
Russia’s VED price registration timeline is independent of the marketing authorisation timeline but must be completed before commercial supply can begin at registered prices. For products that may be included in the VED list, early engagement with the pricing process prevents launch delays.
What Should Companies Check Before Entering the EAEU Market?
Before entering the Eurasian Economic Union (EAEU) pharmaceutical market, companies should evaluate these key regulatory requirements:
Registration Pathway: Determine whether EAEU or an applicable national procedure is appropriate.
CTD Dossier: Verify that the Common Technical Document (CTD) meets applicable EAEU requirements.
Reference State: Identify the reference and concerned member states, where applicable.
Product Information: Review labelling, packaging and product information requirements for target markets.
Pharmacovigilance: Establish the required safety monitoring and reporting systems.
Regulatory Status: Check certificate validity, transitional provisions and pending compliance obligations.
Market Strategy: Assess country-specific requirements, timelines and post-approval responsibilities before planning commercial launch.
How Can DDReg Support EAEU Pharmaceutical Registration?
DDReg supports pharmaceutical companies with EAEU drug registration and regulatory strategy across the EAEU market. Its support can include registration pathway assessment, EAEU CTD and dossier strategy, regulatory intelligence, submission planning, pharmacovigilance, and lifecycle regulatory support.
By combining regional regulatory knowledge with market-specific planning, DDReg helps companies navigate the common EAEU framework while addressing the requirements that remain relevant at the national level.
Conclusion
The EAEU Unified Procedure provides an important mechanism for coordinated pharmaceutical registration across member states, but “unified” does not mean “uniform.” The EAEU pharmaceutical registration process combines common regulatory requirements with member-state responsibilities that companies need to understand before filing.
A successful EAEU market-entry strategy therefore depends on more than preparing an EAEU CTD. Applicants need to select the appropriate registration pathway, understand national requirements, prepare the right registration dossier, and plan pharmacovigilance and post-authorisation activities from the outset. For companies comparing EAEU and EU expansion strategies, understanding these structural differences can help create a more practical and predictable regulatory plan.
Frequently Asked Questions
The EAEU pharmaceutical registration process is the regulatory pathway used to obtain marketing authorisation for medicines within the Eurasian Economic Union. Depending on the product and market strategy, applicants may use the EAEU Unified Procedure or an applicable national pathway. The process generally involves dossier preparation, submission, scientific assessment, responses to regulatory questions, and registration followed by post-authorisation obligations.
An EAEU CTD dossier follows a five-module structure covering administrative information, scientific summaries, quality, nonclinical data, and clinical data. The dossier may also require product information, packaging materials, pharmacovigilance documentation, and other supporting information according to the applicable EAEU requirements.
No. Although the EAEU Unified Procedure and EU MRP/DCP involve coordinated assessment across multiple markets, they operate under different legal and regulatory frameworks. EU MRP generally relies on an existing national marketing authorisation, while DCP is used for simultaneous applications in multiple EU member states. The EAEU Unified Procedure has its own reference-state and participating-state structure.
EAEU registration establishes a common regulatory framework, including EAEU Good Pharmacovigilance Practice requirements, but marketing authorisation holders must also consider how pharmacovigilance requirements are implemented through the national systems and legislation of individual member states. EAEU registration should therefore be accompanied by a clear, market-specific pharmacovigilance strategy.